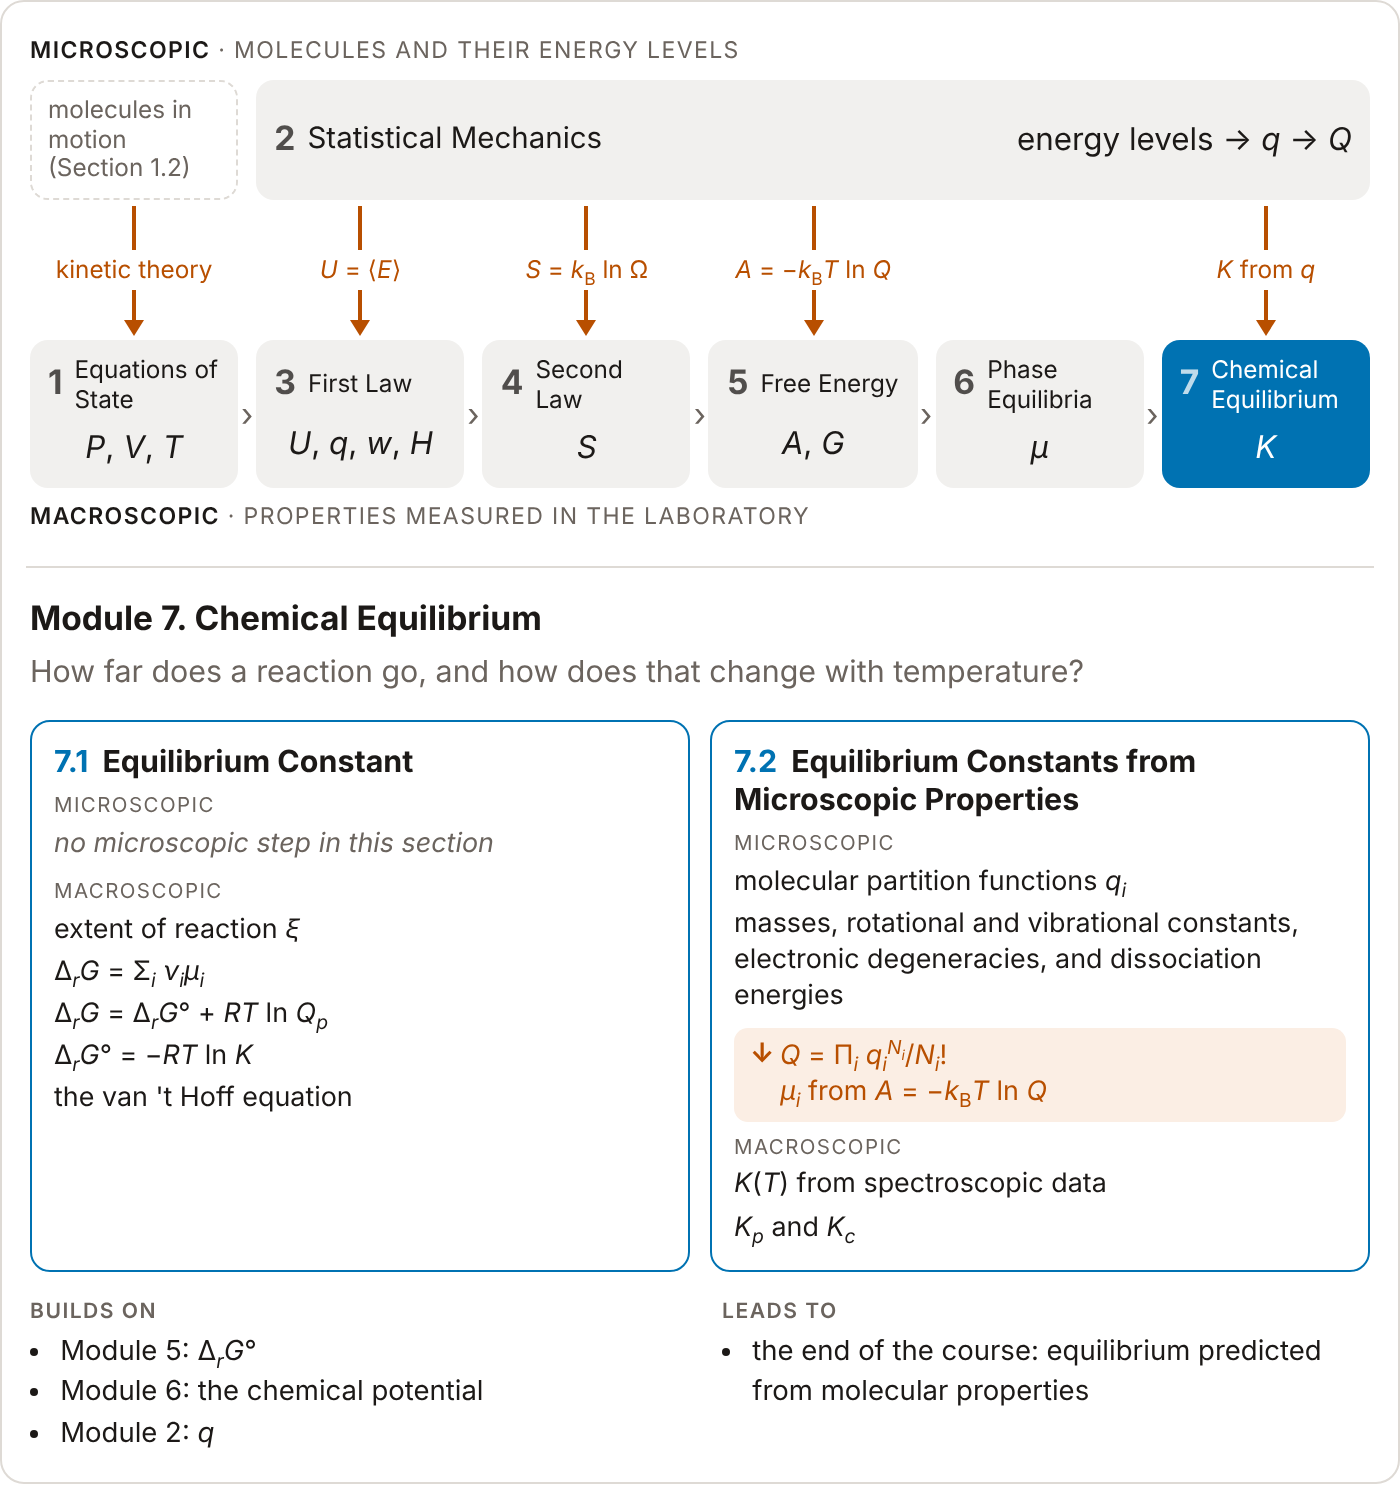
The course map shows where this module sits in the course and which relations connect the microscopic and macroscopic descriptions. Select a module to unfold its sections.
This is the multi-species analog of the phase-equilibrium condition μα=μβ of Section 6.1: instead of moving matter between two phases, matter is redistributed among several chemical species.
Spontaneity: ΔrG<0 drives the reaction forward, ΔrG>0 backward, ΔrG=0 is equilibrium.
Ideal-Gas Chemical Potential and the Standard Pressure
The logarithm contains the ratioPi/P∘, not the pressure itself (Concept Check 2 of Section 7.1). For real gases and solutions, the ratio is replaced by an activity ai.
Reaction Quotient and Equilibrium Constant
Substituting μi=μi∘+RTln(Pi/P∘) into ΔrG=∑iνiμi and grouping terms gives
The first term is linear in ξ (the “bookkeeping” drop in standard chemical potentials); the second is the ideal-mixing term, −TΔmixS, which is zero for pure reactants and for pure products and negative in between.
The minimum of G(ξ) lies where the slope ΔrG=ΔrG∘+RTlnQp is zero. Concept Check 4 of Section 7.1 asks why this point lies between pure reactants and pure products even when ∣ΔrG∘∣≫RT.
Temperature Dependence: the van’t Hoff Equation
Starting from the Gibbs–Helmholtz relation, (∂(G/T)/∂T)P=−H/T2, and ΔrG∘=−RTlnK:
Exothermic reactions (ΔrH∘<0) have K that decreases with T; endothermic reactions have K that increases with T. This is the quantitative form of Le Châtelier’s principle for temperature.
The mathematical structure is identical to the integrated Clausius–Clapeyron equation of Section 6.2, with K in place of the vapor pressure and ΔrH∘ in place of Δhˉvap.
Computing K from Tabulated Thermochemistry
The standard workflow for any gas-phase reaction at a specified T:
Balance the reaction and fix a consistent standard state (P∘=1bar, usually at 298.15 K).
Form ΔrH∘=∑iνiHi∘ and ΔrS∘=∑iνiSi∘ using signed stoichiometry.
Combine into ΔrG∘(T)=ΔrH∘(T)−TΔrS∘(T), and finally K(T)=exp(−ΔrG∘(T)/RT).
Common pitfalls: mixing phases (e.g., H2O(l) vs. H2O(g)), mismatched standard states, and the near-universal unit mismatch between ΔH∘ (kJ/mol) and S∘ (J/mol·K).
Section 7.2: Equilibrium Constants from Microscopic Properties
a condition of the same form as that of Section 7.1. What changes between fixed T,P and fixed T,V is which potential is stationary. The chemical potential is the same quantity in both: μi=(∂G/∂ni)T,P,nj=i=(∂A/∂ni)T,V,nj=i.
Partition Function of an Ideal-Gas Mixture
From factorization over independent subsystems and the indistinguishability correction (Section 2.4),
where qi is the single-molecule partition function for species i. The factorization is exact for an ideal gas because molecules of different species do not interact.
Chemical Potential from A=−kBTlnQ
Using A=−kBTlnQ (Section 4.3) and differentiating with respect to Ni under Stirling’s approximation gives
on a per-mole basis. This is the microscopic counterpart of μi=μi∘(T)+RTln(Pi/P∘) from Section 7.1: the macroscopic expression writes μi in terms of a tabulated μi∘(T) and a partial pressure; the microscopic expression writes the same μi in terms of a counted ratio qi/Ni.
Equilibrium Constant in Terms of Partition Functions
Inserting μi=−RTln(qi/Ni) into ∑iνiμi=0 and exponentiating:
Energy-zero convention: the zero of molecular energy is the dissociated atoms at rest, so the bound v=0 level sits at −D0 and the bond energy enters the electronic factor as e+D0/RT. Vibrational energies are then measured from v=0, which gives qvib=1/(1−e−Θvib/T) in place of the form of Section 2.6. Either zero may be used, provided that qvib and the bond energy refer to the same one (Concept Check 3 of Section 7.2).
With the energy zero at the separated atoms, the zero-point energies are contained in the D0 values, and the vibrational block holds only the “hot-mode” (1−e−Θvib/T) factors.
Physical Reading of the Four Factors at 700 K
Plugging spectroscopic constants for H2, I2, and HI gives K(700K)≈62, to be compared with 56.8 from the NIST-JANAF tables and with a measured value near 54. Agreement within 10 to 15 % is what the rigid-rotor/harmonic-oscillator approximation can be expected to give; the largest single correction, about 4 %, comes from the high-temperature form of qrot for H2.
Translational (≈181): dominated by the mass mismatch between H and I, since mHI2/(mH2mI2)=(mH+mI)2/(4mHmI) is driven far from unity when mI≫mH.
Rotational (≈0.22): I2’s tiny Θrot≈0.05K makes qrot(I2) extremely large, favoring reactants; the symmetry-number prefactor σH2σI2/σHI2=4 nudges toward products but does not overcome this.
Vibrational (≈0.36): the stiff stretches of H2 and HI remain nearly frozen at 700 K, so their factors are within 1 % of unity; the factor is essentially that of I2, whose low-frequency mode is partially excited.
Bond-energy (≈4.3): 2D0HI−D0H2−D0I2≈+8.5kJmol−1 — the two HI bonds together are slightly stronger than one H–H plus one I–I bond.
Every digit of K traces back to a specific molecular property; the relative sizes of the four factors identify which molecular feature most strongly determines the equilibrium.
Consistency with the Macroscopic Route
A student who computes ΔrG∘ from thermochemical tables (Section 7.1) and a student who evaluates K from spectroscopic constants (Section 7.2) arrive at nearly the same number by very different paths. The macroscopic route answers “what is K?” efficiently; the microscopic route answers “why is K what it is?” by decomposing it into separable molecular contributions. Together they close the statistical-mechanical bridge built across Modules 2, 4, 5, and 7.
Applicability: any reaction written with signed stoichiometric coefficients (νi<0 for reactants, νi>0 for products). ξ serves as a single progress variable along the reaction coordinate.
Applicability: any reacting system at equilibrium, at constant T,Por constant T,V. The condition has the same form in both cases; the stationary potential (G vs. A) differs.
D. Ideal-Gas Chemical Potential and Reaction Quotient
Applicability: ideal-gas reactions. Bridges tabulated standard thermochemistry to the equilibrium constant and to reaction direction via the comparison Qp≶K.
F. van’t Hoff Equation (differential and integrated forms)
Applicability: temperature dependence of K; integrated form assumes ΔrH∘ is approximately constant on [T1,T2]. Structurally identical to the integrated Clausius–Clapeyron equation of Section 6.2.
Applicability: ideal gases (non-interacting species); follows from factorization over independent subsystems with the indistinguishability correction (Section 2.4).
Applicability: ideal-gas species in the thermodynamic limit, where Stirling’s approximation is valid. Microscopic counterpart of the macroscopic expression in (D).
Applicability: diatomic ideal gas in the rigid-rotor/harmonic-oscillator approximation (Section 2.8), with the energy zero chosen at the dissociated atoms at rest, so that vibrational energies are measured from v=0. σ=2 for homonuclear diatomics, σ=1 for heteronuclear.
Applicability: the specific Δν=0 benchmark reaction in the rigid-rotor/harmonic-oscillator approximation. Separates K into translational (mass), rotational (symmetry and Θrot), vibrational (hot-mode), and bond-energy factors — each traceable to identifiable molecular structure.
Model air as 4.00mol of N2 and 1.00mol of O2. The table lists data for NO(g) from the NIST-JANAF tables. Use R=8.3145Jmol−1K−1.
T (K)
ΔfG∘ (kJmol−1)
ΔfH∘ (kJmol−1)
1000
77.775
90.437
2000
65.060
90.494
(a) Find ΔrG∘ and K at 2000K.
(b) Write K in terms of the extent of reaction at equilibrium, ξeq. Find ξeq and the mole fraction of NO at 2000K.
(c) Estimate K at 2500K with the van’t Hoff equation and ΔrH∘ at 2000K. Compare the estimate with K=3.52×10⁻³ from the same tables.
(d) The equilibrium mixture of part (b) is cooled to 1000K so quickly that its composition does not change. Find Qp, K, and ΔrG at 1000K. In which direction is the reaction spontaneous?
Solution
ΔrG∘ and K at 2000K
N2 and O2 are elements in their reference states, so ΔfG∘=0 for both and
Qp>K and ΔrG>0, so the reverse reaction is spontaneous: nitric oxide should decompose into N2 and O2. This result gives the direction of the change and says nothing about its rate.
Result.K=4.00×10⁻⁴ at 2000K, with ξeq=0.0197mol and yNO=0.0079. The van’t Hoff equation gives K=3.52×10⁻³ at 2500K. At 1000K the frozen mixture has Qp=4.00×10⁻⁴>K=7.5×10⁻⁹ and ΔrG=+90.49kJmol−1, so the reverse reaction is spontaneous.
Problem 2. Pressure and the extent of a dissociation
At 700K the NIST-JANAF tables give ΔfG∘=−187.054kJmol−1 for COCl2(g) and −173.518kJmol−1 for CO(g). A vessel initially holds 1.00mol of phosgene and no products. Use R=8.3145Jmol−1K−1 and P∘=1bar.
(a) Find ΔrG∘ and K at 700K.
(b) Write K in terms of ξeq and the total pressure P, and solve the expression for ξeq.
(c) Find ξeq and the three partial pressures at P=1.00bar and at P=0.100bar.
Solution
ΔrG∘ and K
Cl2(g) is an element in its reference state, so its ΔfG∘ is zero.
As a check, (0.230)2/0.540=0.098 and (0.0413)2/0.0174=0.098: both mixtures have Qp=K. The dissociation has Δν=+1, and a tenfold drop in pressure raises ξeq from 0.298 to 0.703mol while K stays the same.
Result.ΔrG∘=+13.536kJmol−1 and K=0.0977 at 700K. ξeq=K/(K+P/P∘), which is 0.298mol at 1.00bar and 0.703mol at 0.100bar.
Problem 3. The Gibbs energy minimum of hot carbon dioxide
At this temperature the NIST-JANAF tables give ΔfG∘=−395.461kJmol−1 for CO2(g) and −367.826kJmol−1 for CO(g). The system starts as 1.00mol of CO2 and stays at 3000K and P=P∘=1bar. Use R=8.3145Jmol−1K−1.
(a) Find ΔrG∘ and K.
(b) Write the amounts ni(ξ) and the mole fractions yi(ξ). Then write G(ξ)−G(0) in the form used for NO2 in Section 7.1.
(c) Write Qp as a function of ξ, and confirm that ξeq=0.438mol satisfies Qp=K.
(d) Evaluate G(ξ)−G(0) at ξeq and at ξ=1mol. Evaluate the slope ΔrG at ξ=0.200mol and at ξ=0.700mol.
The slope is ΔrG=ΔrG∘+RTlnQp, with Qp from step 3.
ξ (mol)
Qp
RTlnQp (kJmol−1)
ΔrG (kJmol−1)
0.200
0.0754
−64.48
−36.85
0.438
0.3303
−27.63
0.00
0.700
1.188
+4.30
+31.93
The slope is negative at ξ=0.200mol, zero at ξeq, and positive at ξ=0.700mol.
Result.K=0.3303 and ξeq=0.438mol. Relative to pure CO2, G is −19.31kJ at ξeq and +3.82kJ at ξ=1mol. ΔrG is −36.85kJmol−1 at ξ=0.200mol and +31.93kJmol−1 at ξ=0.700mol.
Problem 4. Dissociation of iodine from molecular constants
The table lists the constants of I2 from Section 7.2 and those of the atom, whose ground level has the degeneracy g1=4.
Species
M (gmol−1)
Θrot (K)
Θvib (K)
D0 (kJmol−1)
σ
g1
I2
253.808
0.05377
309
148.81
2
1
I
126.904
4
Take the zero of energy at the separated atoms, as in Section 7.2, so that qI=(V/ΛI3)g1. Use h=6.6261×10⁻³⁴Js, kB=1.3806×10⁻²³JK−1, NA=6.0221×10²³mol−1, R=8.3145Jmol−1K−1, and P∘=1bar.
(a) Find q/V for the atom and for the molecule at 1000K.
(b) Find Kc and Kp at 1000K.
(c) Find ΔrG∘ from Kp. Compare it with the NIST-JANAF value ΔfG∘(I,g)=24.039kJmol−1 at 1000K, for which the reference state of iodine is I2(g).
(d) One mole of I2 comes to equilibrium at 1000K and a total pressure of 1.00bar. Find ξeq and the two partial pressures.
Solution
The atom
The mass of one atom is mI=M/NA=2.107×10−25kg, so
The molecule has twice the mass, so ΛI2=ΛI/2=3.465×10−12m and 1/ΛI23=2.403×1034m−3. The other three factors of the diatomic partition function are
The tables give ΔrG∘=2(24.039)=48.08kJmol−1, which corresponds to K=3.08×10⁻³. The model value of ΔrG∘ is about 0.3kJmol−1 lower, and its Kp is 4% higher.
Composition at 1.00bar
The amounts are nI2=1−ξ and nI=2ξ, with ntot=1+ξ. Then
Result. At 1000K, qI/V=3.398×1034m−3 and qI2/V=4.982×1046m−3, which give Kc=2.318×10²²m−3, Kp=3.20×10⁻³, and ΔrG∘=+47.76kJmol−1 (NIST-JANAF: 48.08kJmol−1). At 1.00bar, ξeq=0.0283mol, PI=0.0550bar, and PI2=0.945bar.
Hill (1986), Chapter 10 (chemical equilibrium in ideal gas mixtures).
Original papers
Hoff (1884) contains the relation between the equilibrium constant and the temperature, the van’t Hoff equation of Section 7.1. The English edition, translated by T. Ewan, is Studies in Chemical Dynamics (1896).
McQuarrie, D. A., & Simon, J. D. (1997). Physical Chemistry: A Molecular Approach. University Science Books.
Dill, K. A., & Bromberg, S. (2011). Molecular Driving Forces: Statistical Thermodynamics in Biology, Chemistry, Physics, and Nanoscience (2nd ed.). Garland Science.
Hill, T. L. (1986). An Introduction to Statistical Thermodynamics. Dover Publications.
van ’t Hoff, J. H. (1884). Études de dynamique chimique. Frederik Muller.
Le Chatelier, H. (1884). Sur un énoncé général des lois des équilibres chimiques. Comptes Rendus Hebdomadaires Des Séances de l’Académie Des Sciences, 99, 786–789.
Taylor, A. H., & Crist, R. H. (1941). Rate and Equilibrium Studies on the Thermal Reaction of Hydrogen and Iodine. Journal of the American Chemical Society, 63(5), 1377–1385. 10.1021/ja01850a071