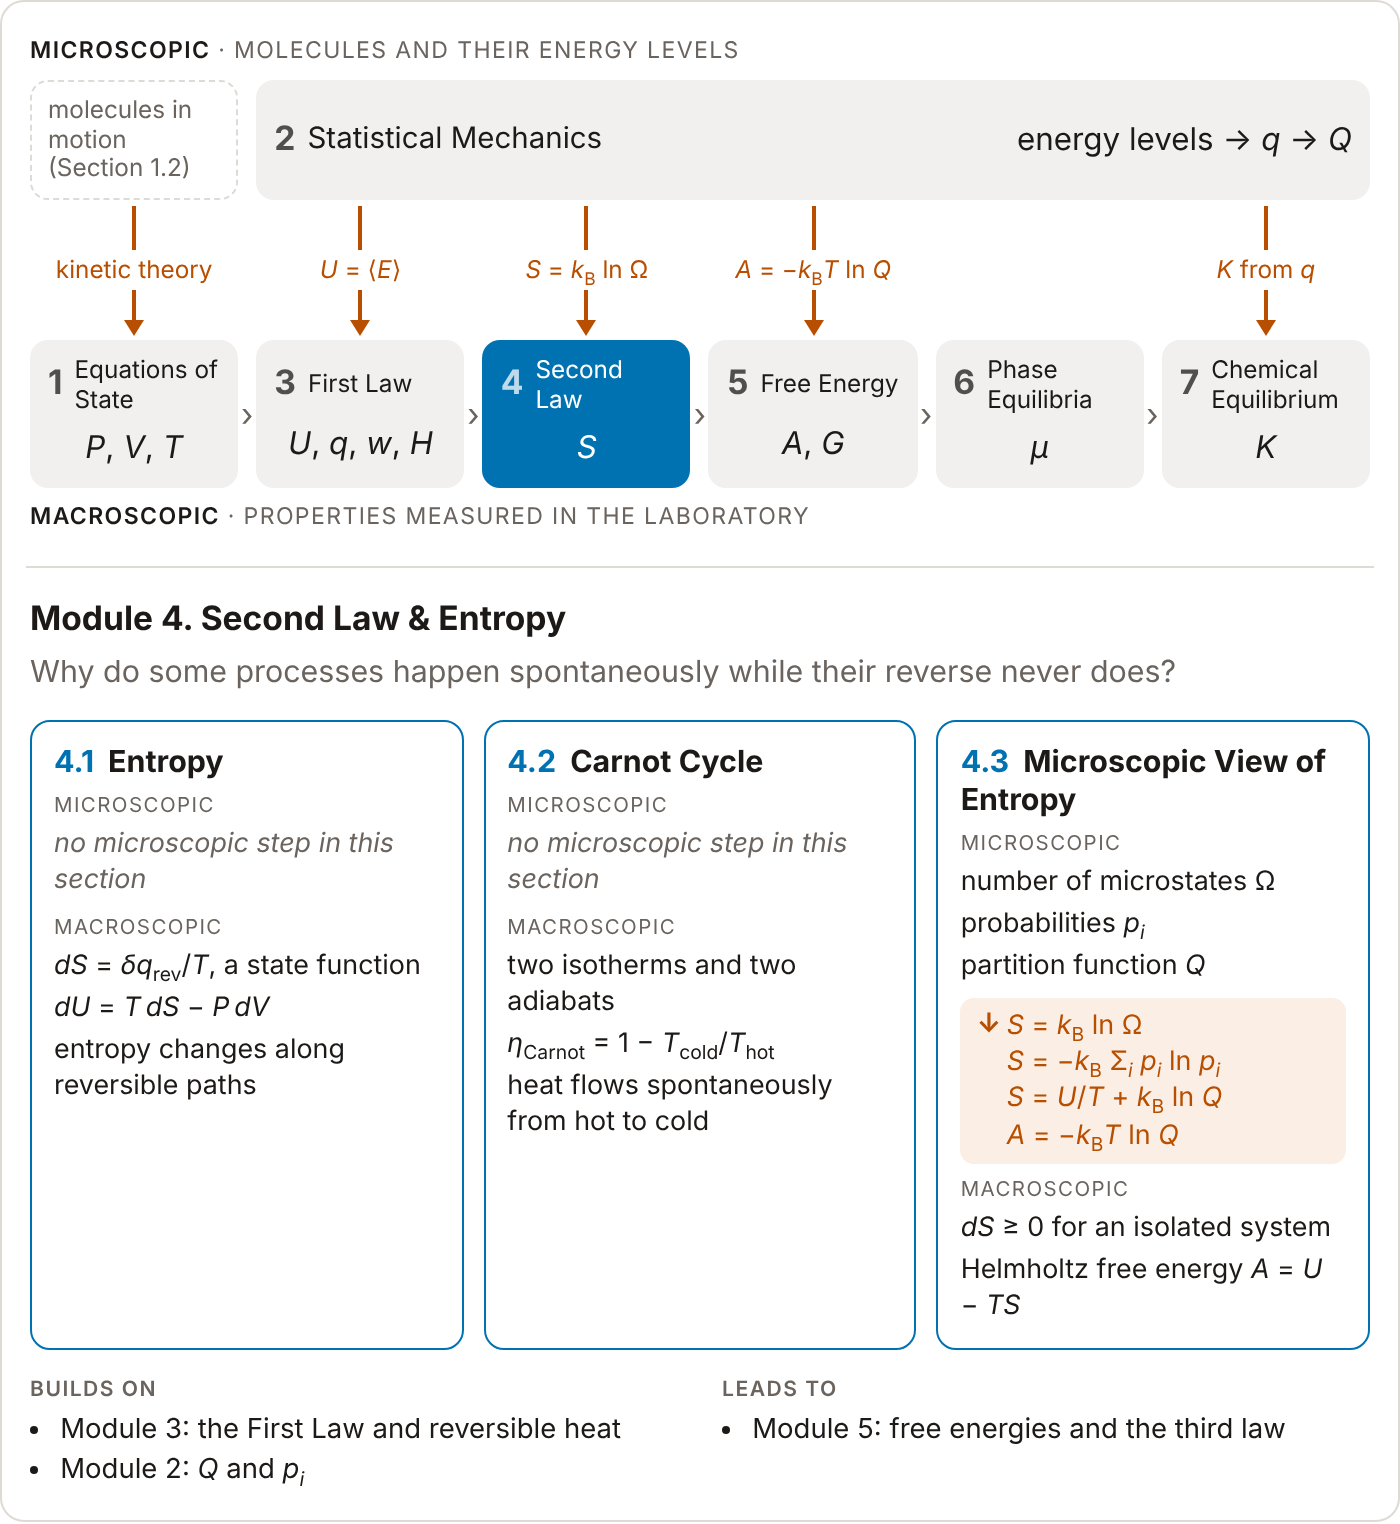
The course map shows where this module sits in the course and which relations connect the microscopic and macroscopic descriptions. Select a module to unfold its sections.
1. Checklist of Key Concepts
Section 4.1: Entropy
See Section 4.1.
Exothermicity Does Not Guarantee Spontaneity
(exothermic) often favors spontaneity but does not determine it. Counter-example: mixing of two ideal gases is spontaneous even though .
A complete account of directionality requires entropy.
Definition of Entropy
Entropy is a state function defined by
The finite change between states and is , evaluated along any reversible path connecting those states.
Exactness of
is inexact, but dividing by produces an exact differential. Verification for an ideal gas: the cross-derivative test on and gives matching mixed partials (both zero).
Entropy of Irreversible Processes
Because is a state function, depends only on the endpoints. To compute for an irreversible process, find any convenient reversible path between the same initial and final states and integrate along that path.
Fundamental Thermodynamic Relation
Combining and in the First Law:
The natural variables of are and . This relation replaces two inexact differentials (, ) with three exact ones (, , ).
The relation is derived for a reversible process and holds for any change between equilibrium states, because it contains only state functions.
Two Expansions Between the Same States (Ideal Gas)
Reversible isothermal expansion: .
Free expansion into vacuum: and , with the same , because is a state function.
Section 4.2: Carnot Cycle
See Section 4.2.
Structure of the Carnot Cycle
A fully reversible cycle operating between a hot reservoir () and a cold reservoir (), consisting of four steps:
: reversible isothermal expansion at (system absorbs heat ).
: reversible adiabatic expansion (system cools, ).
: reversible isothermal compression at (system releases heat ).
: reversible adiabatic compression (system warms, ).
Entropy Bookkeeping Around the Cycle
Adiabatic steps are isentropic: .
Isothermal steps carry entropy: , .
Over one complete cycle, (state function returns to initial value), so .
Carnot Efficiency
The efficiency of an engine is the fraction of the absorbed heat that becomes net work output: .
For the Carnot cycle:
This is the maximum possible efficiency for any engine operating between two temperatures. It depends only on the reservoir temperatures.
Second Law for an Isolated System
Spontaneous change requires , with equality for a reversible change.
Direction of Spontaneous Heat Flow
Two subsystems at different temperatures inside an insulated boundary: .
For , heat must flow from hot to cold ( when ).
Section 4.3: Microscopic View of Entropy
See Section 4.3.
Entropy Increases Until Equilibrium (Isolated Systems)
The second law for an isolated system: , with equality at equilibrium.
Entropy rises during spontaneous processes and reaches its maximum at equilibrium.
The Clausius Inequality
Equality for reversible processes; strict inequality for irreversible processes.
is the temperature at which the heat is transferred to the system (for a reservoir, the temperature of the reservoir).
Irreversibility produces additional entropy beyond the entropy carried by heat flow.
For an isolated system (): reduces to .
Summed over one cycle of any engine between two reservoirs, it gives .
Boltzmann’s Formula (Microcanonical Ensemble)
where is the number of accessible microstates at fixed , , . The logarithm ensures entropy is extensive (additive for independent subsystems).
Gibbs Entropy (General Probability Distribution)
Reduces to Boltzmann’s formula when all microstates are equally probable ().
Applies to the canonical ensemble () and any other ensemble.
Canonical Entropy Identity
Evaluating the Gibbs entropy with canonical probabilities yields
Directly ties the macroscopic state function to the partition function from Module 2.
Helmholtz Free Energy
Rearranging the canonical entropy identity:
Natural variables of are and .
All canonical thermodynamic quantities (, , , ) can be derived from by differentiation.
Defining is the same Legendre-transform strategy used to define in Section 3.3: replace a hard-to-control variable () with an easy-to-control one ().
Microscopic Consistency
For a reversible process, the Gibbs entropy with canonical probabilities gives , the definition of entropy in Section 4.1.
This is consistent with the microscopic First Law from Section 3.2: and .
The restriction to reversible processes matters: in a free expansion the entropy increases although no heat flows.
2. Checklist of Most Important Equations
Below is a unified list of the major equations from Sections 4.1–4.3.
A. Entropy Definition
Applicability: any system. The integral must be evaluated along a reversible path, but itself depends only on the endpoints.
B. Fundamental Thermodynamic Relation
Applicability: simple closed system with -only work, for any change between equilibrium states. Natural variables of are and .
C. Entropy Change for an Ideal Gas (from and )
For a monatomic ideal gas with constant : .
D. Carnot Efficiency
Applicability: maximum efficiency for any heat engine operating between two reservoir temperatures. The efficiency is defined as .
E. Clausius Inequality
Equality for reversible processes; strict inequality for irreversible processes. is the temperature at which the heat is transferred.
For isolated systems (): .
F. Boltzmann Entropy
Applicability: isolated systems (microcanonical ensemble) where all accessible microstates are equally probable.
G. Gibbs Entropy
Applicability: any probability distribution over microstates. Reduces to Boltzmann’s formula in the microcanonical limit.
H. Canonical Entropy Identity
Applicability: canonical ensemble (fixed , , ). Connects macroscopic entropy to the partition function.
I. Helmholtz Free Energy
Applicability: canonical ensemble. Natural variables are and . All other canonical thermodynamic quantities can be obtained by differentiation.
3. Practice Problems
Each problem is a multi-part question of exam length. Work through it on paper before you open the solution.
Problem 1. Three reversible paths, one 
Draws on Section 4.1.
One mole of oxygen, treated as an ideal gas with , is taken from state A (, ) to state B (, ). Three reversible paths connect the states:
Path 1. The gas is heated at constant volume to and then expanded isothermally at to .
Path 2. The gas is expanded isothermally at to and then heated at constant volume to .
Path 3. The gas is compressed adiabatically until its temperature is and then expanded isothermally at to .
Use . Along a reversible adiabat of this gas, is constant (Section 3.2).
(a) Find and for each step of Path 1.
(b) Find and for each step of Path 2.
(c) Add the steps to obtain and for Paths 1 and 2. Compare with the entropy change of an ideal gas from Section 4.1.
(d) Find the volume at the end of the adiabatic step of Path 3. Then find and for Path 3.
Solution
Relations for the two kinds of step
The heat capacity is .
At constant volume, , so
At constant temperature, (Example 1 of Section 4.1), so
Path 1, heating at constant volume ()
Path 1, isothermal expansion at
Path 2, isothermal expansion at
The volume ratio is the same as in step 3, so , and
Path 2, heating at constant volume ()
The heat capacity of an ideal gas does not depend on the volume, so this step has the values of step 2: and .
Totals for Paths 1 and 2
Path (J) () Path 1 Path 2 For constant , the entropy change of an ideal gas from Section 4.1 is
which contains only the temperatures and volumes of the two states.
Path 3, adiabatic compression
With , the volume at the end of the adiabatic step is
On a reversible adiabat , so and for this step.
Path 3, isothermal expansion at
Result. The three paths have different heats and the same entropy change.
| Path | (J) | () |
|---|---|---|
| Path 1 | 7335 | 18.52 |
| Path 2 | 5952 | 18.52 |
| Path 3 | 7778 | 18.52 |
Problem 2. The four steps of a Carnot cycle
Draws on Section 4.2.
A Carnot engine uses one mole of a diatomic ideal gas with as its working substance. The reservoirs are at and . The cycle has the four reversible steps of Section 4.2: isothermal expansion at , adiabatic expansion , isothermal compression at , and adiabatic compression . The first two volumes are and .
Use . Along a reversible adiabat of this gas, is constant (Section 3.2).
(a) Find and .
(b) Find and for each of the four steps.
(c) Find the net work output and the efficiency from the heats of part (b). Compare the efficiency with .
(d) Find of the gas for each step and for the whole cycle.
Solution
Volumes at the ends of the adiabats
For step , , so
Step connects the same two temperatures, so
The two isotherms therefore have reciprocal volume ratios: .
Isothermal steps
For an ideal gas at constant temperature , so .
The work is and .
Adiabatic steps
With , , and .
Net work output and efficiency
The work of the two adiabatic steps cancels. In the notation of Section 4.2,
Entropy changes of the gas
On the isotherms, :
On the reversible adiabats, . The sum over the cycle is .
Result. and . The engine delivers per cycle with , equal to .
| Step | (J) | (J) | (J) | () |
|---|---|---|---|---|
| +4571 | −4571 | 0 | +7.618 | |
| 0 | −4157 | −4157 | 0 | |
| −3047 | +3047 | 0 | −7.618 | |
| 0 | +4157 | +4157 | 0 | |
| Cycle | +1524 | −1524 | 0 | 0 |
Problem 3. Heat flow between two bodies
Draws on Section 4.1 and Section 4.2.
A copper block of mass at is dropped into of water at inside an insulated container. The specific heat capacities are for copper and for water; treat them as constant. Neglect the thermal expansion of the copper and the water, so that the two bodies exchange only heat with each other, and neglect the heat capacity of the container.
(a) Find the final temperature.
(b) Find of the copper and of the water.
(c) Find the total entropy change, and compare its sign with the Second Law for an isolated system in Section 4.2.
Solution
Heat capacities
Final temperature
The container is insulated and no work is done, so the internal energy of the copper and the water together is constant:
Solving for ,
The copper gives up of heat to the water.
Entropy change of each body
The flow of heat between the two bodies is irreversible. Entropy is a state function, so of each body is found on a reversible path between its initial and final states, on which the body is heated or cooled slowly. With no work, , and
Carrying one more figure in the final temperature, ,
Total entropy change
The copper and the water together form an isolated system, and the total is positive, in agreement with for a spontaneous change in an isolated system.
Result. . The copper loses of entropy, the water gains , and the total entropy increases by .
Problem 4. Electronic entropy of an ion with one electron
Draws on Section 4.3 and Section 2.2.
The ion has a single electron. Without spin–orbit coupling, its ten lowest states (five orbitals, each with two spin orientations) have the same energy. Spin–orbit coupling splits them into a lower level of four states and an upper level of six states, which lies above the lower level (NIST Atomic Spectra Database). The next level lies at and is left out of this problem.
Treat one free ion at as the system, so that is a sum over its ten states. Use , and give each entropy in units of .
(a) Find the entropy if the ten states have the same energy.
(b) With the splitting, find and the probability of each state of the lower level and of the upper level.
(c) Find the Gibbs entropy from the probabilities of part (b).
(d) Use to find the entropy a second way, with measured from the lower level.
Solution
Ten states of equal energy
The ten states are equally likely, , so the Gibbs entropy reduces to Boltzmann’s formula with :
The splitting in units of
Let be the energy of the upper level above the lower level.
Partition function and probabilities
Four states have the Boltzmann factor 1 and six states have .
Each state of the lower level has probability , and each state of the upper level has probability :
The lower level holds of the probability and the upper level holds .
Gibbs entropy
Entropy from the partition function
Only the six upper states contribute to the energy, so and
Then
Result. for ten states of equal energy. With the spin–orbit splitting, , , , and at by both routes.
4. Further Reading
Textbooks
McQuarrie & Simon (1997), Chapter 20 (entropy and the second law of thermodynamics).
Dill & Bromberg (2011), Chapter 5 (entropy and the Boltzmann law) and Chapter 6 (thermodynamic driving forces).
Hill (1986), Chapter 2 (the second and third laws in terms of ensembles).
Original papers
Carnot (1824) analyzes the ideal heat engine of Section 4.2 and shows that its efficiency depends only on the temperatures of the two reservoirs. The English translation by R. H. Thurston is Reflections on the Motive Power of Heat (Wiley, New York, 1897).
Clausius (1865) introduces the word entropy for the state function of Section 4.1.
Sharp & Matschinsky (2015) translate into English the paper of 1877 in which Boltzmann relates the entropy to the number of ways of distributing the energy among the molecules (Section 4.3).
- McQuarrie, D. A., & Simon, J. D. (1997). Physical Chemistry: A Molecular Approach. University Science Books.
- Dill, K. A., & Bromberg, S. (2011). Molecular Driving Forces: Statistical Thermodynamics in Biology, Chemistry, Physics, and Nanoscience (2nd ed.). Garland Science.
- Hill, T. L. (1986). An Introduction to Statistical Thermodynamics. Dover Publications.
- Carnot, S. (1824). Réflexions sur la puissance motrice du feu et sur les machines propres à développer cette puissance. Bachelier.
- Clausius, R. (1865). Ueber verschiedene für die Anwendung bequeme Formen der Hauptgleichungen der mechanischen Wärmetheorie. Annalen Der Physik, 201(7), 353–400. 10.1002/andp.18652010702
- Sharp, K., & Matschinsky, F. (2015). Translation of Ludwig Boltzmann’s Paper “On the Relationship between the Second Fundamental Theorem of the Mechanical Theory of Heat and Probability Calculations Regarding the Conditions for Thermal Equilibrium” (Wien. Ber. 1877, 76, 373–435). Entropy, 17(4), 1971–2009. 10.3390/e17041971