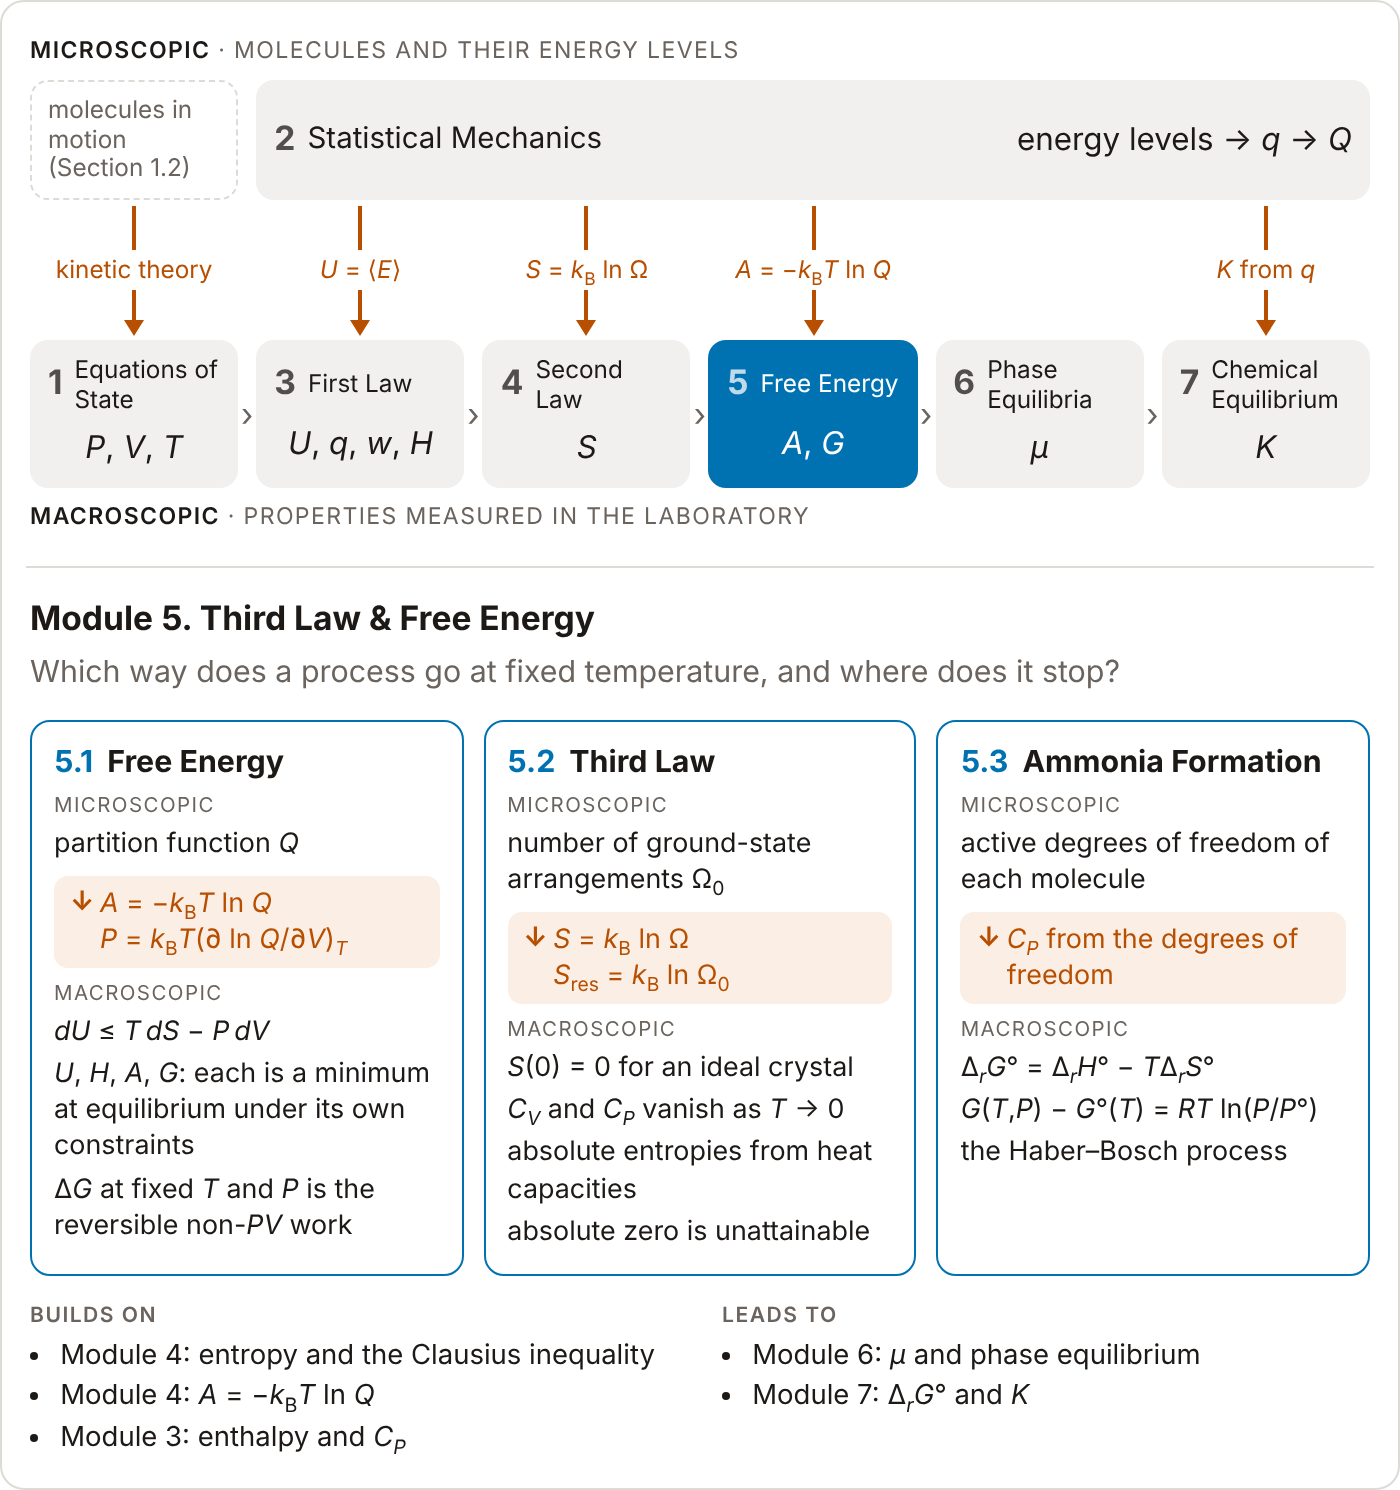
The course map shows where this module sits in the course and which relations connect the microscopic and macroscopic descriptions. Select a module to unfold its sections.
For any change between equilibrium states of a closed system with PV-only work, dU=TdS−PdV (Section 4.1).
For a closed system with PV-only work that is in thermal and mechanical equilibrium with its surroundings, but not necessarily in equilibrium internally, the First Law and the Clausius inequality dS≥δq/T give
with equality when every change in the system is reversible. The strict inequality holds when an irreversible process takes place inside the system, and it gives the direction of spontaneous change.
Extremum Principles for Thermodynamic Potentials
Under different experimental constraints, a particular potential is minimized at equilibrium:
Constant S,V: dU≤0 ⇒ U minimized.
Constant S,P: dH≤0 ⇒ H minimized.
Constant T,V: dA≤0 ⇒ A minimized.
Constant T,P: dG≤0 ⇒ G minimized.
The T,V and T,P cases cover the vast majority of laboratory conditions.
Legendre Transforms and Natural Variables
Each potential is obtained from U by a Legendre transform that swaps an extensive variable for its conjugate intensive variable, matching the natural variables to the experimentally controllable ones.
Summary of natural variables and equilibrium differentials:
potential
definition
differential
natural variables
U
—
dU=TdS−PdV
S,V
H
U+PV
dH=TdS+VdP
S,P
A
U−TS
dA=−SdT−PdV
T,V
G
U+PV−TS
dG=−SdT+VdP
T,P
Defining A and G extends the strategy used in Section 3.3 to define H.
Conjugate Variables as Derivatives
At equilibrium, intensive variables are read off as slopes of a potential along its natural-variable axes. For example:
Differentiating yields the Sackur–Tetrode entropyS=NkB[ln(V/NΛ3)+5/2], the equation of state PV=NkBT, and the internal energy U=23NkBT — all from a single function.
Gibbs Free Energy and Non-PV Work
When non-PV work is present, the First Law gains a δwnon-PV term. At constant T and P:
As T→0K, the entropy of any pure crystalline substance approaches a constant. For an ideal crystal (no defects, impurities, or disorder), that constant is zero:
This fixes the otherwise-undetermined integration constant in ΔS=∫δqrev/T.
Statistical-Mechanical Justification
From Boltzmann’s formula S=kBlnΩ (Section 4.3), a unique ground state has Ω0=1, so S(0)=kBln1=0. The third law is the statement that a system with a nondegenerate ground state has zero entropy at absolute zero.
Residual Entropy
If the ground state is degenerate or the system has frozen-in disorder (Ω0>1), entropy approaches a nonzero constant:
Solid CO (two nearly isoenergetic orientations per molecule): Ω0=2N, giving Sres,m=Rln2≈5.76Jmol−1K−1 (experiment: ≈4.6, indicating partial ordering).
Ice (Pauling’s model with ice rules): Ω0≈(3/2)N, giving Sres,m=Rln(3/2)≈3.37Jmol−1K−1 (experiment: ≈3.41).
Heat Capacities Must Vanish as T→0
For the integral S(T)=S(0)+∫0T(CP/T′)dT′ to converge, CP (and similarly CV) must go to zero at T=0.
This requirement can be compared with the classical (equipartition) heat capacity and with the Einstein heat capacity of Section 2.6.
Unattainability of Absolute Zero (Nernst form)
No finite sequence of thermodynamic operations can cool a system to exactly T=0. Each successive cooling step removes less entropy than the previous one; the process converges but never reaches zero.
Absolute (Third-Law) Entropies
With S(0)=0 fixed for ideal crystals, absolute entropies are computed by integrating CP/T from 0 to T and adding phase-transition contributions:
ΔG determines which direction is favored at equilibrium; activation barriers determine how fast equilibrium is reached. These are independent questions.
For N2+3H2→2NH3 at 298.15 K: ΔrH∘≈−92kJmol−1, ΔrS∘≈−198JK−1mol−1, ΔrG∘≈−33kJmol−1. Thermodynamically spontaneous but kinetically inert because of the N≡N triple bond.
A catalyst lowers the activation barrier without changing ΔrG∘ or the equilibrium constant — it accelerates forward and reverse reactions equally.
Temperature Effect for ΔrS∘<0
From ΔrG∘=ΔrH∘−TΔrS∘, a negative ΔrS∘ means −TΔrS∘>0 grows with T, driving ΔrG∘ upward.
At high enough T, ΔrG∘ changes sign and the reaction becomes non-spontaneous at 1 bar.
At 500 °C: P≈320bar. At 400 °C: P≈86bar. The Haber–Bosch operating window (400–500 °C, 150–350 bar) is the engineering compromise predicted by these estimates.
Engineering Knobs Summary
Catalyst: speeds up both directions; does not change ΔrG∘ or K.
Temperature: raises rate (Arrhenius) but pushes ΔrG∘ upward when ΔrS∘<0.
Pressure: compensates for the temperature penalty when Δν<0.
Applicability: closed system with PV-only work, in thermal and mechanical equilibrium with its surroundings. Equality when every change in the system is reversible; strict inequality when an irreversible process takes place inside the system.
B. Thermodynamic Potentials and Their Differentials (fixed N, PV-only work)
Applicability: reaction data from a reference temperature Ti to a target temperature Tf, assuming temperature-independent heat capacities over the interval.
where Λ is the thermal wavelength, which depends only on T, and ϕ(T) collects the contributions of the rotations and vibrations of the molecules, which also depend only on T. For carbon dioxide, am=3.6551L2barmol−2 and bm=0.042816Lmol−1 (the values used in Section 1.4). Use R=8.3145Jmol−1K−1 and 1Lbar=100J.
(a) Use P=−(∂A/∂V)T to find the equation of state.
(b) The fluid expands isothermally from V1 to V2. Find expressions for ΔA, for ΔS from S=−(∂A/∂T)V, and for ΔU.
(c) Evaluate ΔA, ΔS, and ΔU for one mole of carbon dioxide that expands from 0.300L to 1.00L at 320K.
(d) At constant temperature, ΔA is the reversible work (Section 5.1). Compare the reversible work of part (c) with that of an ideal gas between the same volumes.
The van der Waals fluid has wrev=ΔA=−2644J. The term with bm changes the logarithmic part from −3203J to −3497J, and the term with am adds +853J.
Result.A returns the van der Waals equation of state. For the expansion of carbon dioxide, ΔS=10.93JK−1, ΔU=+853J, and ΔA=wrev=−2644J, against −3203J for an ideal gas.
Problem 2. Standard entropy of oxygen, mode by mode
The tables of Sections 2.6 to 2.8 give, for O2, Θrot=2.080K with σ=2, Θvib=2274K, and a ground electronic level of degeneracy g1=3 whose first excited level is too high to be populated at room temperature. The molar mass is 31.998gmol−1. Treat O2 as an ideal gas at T=298.15K and P=1bar.
Use h=6.62607×10⁻³⁴Js, kB=1.380649×10⁻²³JK−1, NA=6.02214×10²³mol−1, and R=8.3145Jmol−1K−1.
(a) Find the translational entropy per mole from the Sackur–Tetrode equation.
(b) For each of the other modes, the molar entropy is Sm=Um/T+Rlnq, with Um and q of that mode. Find the rotational entropy with qrot=T/(σΘrot) and Um=RT.
(c) Find the vibrational entropy and the electronic entropy.
(d) Add the four contributions, and compare the total with S∘(298.15K)=205.147Jmol−1K−1 from the NIST-JANAF tables.
Solution
Thermal wavelength and volume per molecule
The mass of one molecule is m=(0.031998kgmol−1)/NA=5.3134×10−26kg.
Suppose that the molecules of a crystal freeze into their orientations independently of one another.
In solid nitrous oxide, the linear molecule NNO can lie in either of two directions (NNO or ONN).
In solid perchloryl fluoride, FClO3, the chlorine atom is surrounded by one fluorine atom and three oxygen atoms at the corners of a tetrahedron, and the fluorine atom can occupy any of the four corners.
Use R=8.3145Jmol−1K−1.
(a) Find Ω0 for N molecules and the molar residual entropy of each crystal if all orientations are equally likely.
(b) Suppose instead that the two orientations of a linear molecule have probabilities p and 1−p. Use the Gibbs entropy to find the molar residual entropy as a function of p, and evaluate it for p=0.50 and p=0.75.
(c)Section 5.2 quotes a measured residual entropy of about 4.6Jmol−1K−1 for CO. Find, to two figures, the value of p at which the model of part (b) gives this entropy.
Solution
Equally likely orientations
Each molecule has ω orientations, and the molecules are independent, so Ω0=ωN and
For one molecule with two orientations, the Gibbs entropy is −kB[plnp+(1−p)ln(1−p)]. The entropies of independent molecules add (Section 4.3), so for one mole
The equation −R[plnp+(1−p)ln(1−p)]=4.6Jmol−1K−1 has no solution in closed form. The entropy falls as p rises above 0.50, and step 2 shows that p is slightly above 0.75:
p
Sres,m (Jmol−1K−1)
0.75
4.68
0.76
4.58
0.77
4.48
The measured value lies between the first two rows, closer to the second, so p≈0.76.
Result.Sres,m=Rln2=5.763Jmol−1K−1 for two orientations and Rln4=11.526Jmol−1K−1 for four. With unequal probabilities, Sres,m=−R[plnp+(1−p)ln(1−p)], which is 4.68Jmol−1K−1 at p=0.75 and matches the measured value for CO at p≈0.76. This model keeps the molecules independent; Section 5.2 describes the same shortfall through correlations between neighboring molecules.
Problem 4. When does steam reforming become spontaneous?
(b) Treat ΔrH∘ and ΔrS∘ as independent of temperature, and find the temperature at which ΔrG∘=0.
(c) Find ΔrG∘ at 1200K with the constant heat capacities of Section 5.3: CP=27R for the linear molecules and CP=4R for the nonlinear molecules. Compare the result with −77.92kJmol−1 from the NIST-JANAF tables.
(d) Reformers operate at about 25bar. Find ΔrG at 1200K with every species at 25bar, starting from your result of part (c).
The NIST-JANAF value is −77.92kJmol−1. The estimate has the right sign and is 11.8kJmol−1 too negative. The model leaves out the vibrations: in the same tables the heat capacity of methane at 1200K is 78.8Jmol−1K−1, more than twice 4R.
Pressure
The reaction turns two moles of gas into four, so Δν=(1+3)−(1+1)=+2. With every species at P=25bar,
Result. At 298.15K, ΔrH∘=+206.17kJmol−1, ΔrS∘=+214.61Jmol−1K−1, and ΔrG∘=+142.19kJmol−1. ΔrG∘ changes sign near 961K. At 1200K the constant-CP model gives ΔrG∘=−89.7kJmol−1 (tabulated: −77.9kJmol−1), and ΔrG=−25.5kJmol−1 with every species at 25bar.
McQuarrie & Simon (1997), Chapter 21 (entropy and the third law of thermodynamics) and Chapter 22, Sections 22.1 to 22.7 (Helmholtz and Gibbs energies).
Dill & Bromberg (2011), Chapter 8 (laboratory conditions and free energies) and Chapter 9 (Maxwell’s relations and mixtures).
Hill (1986), Chapter 2 (the second and third laws in terms of ensembles).
Original papers
Clayton & Giauque (1932) measure the entropy of carbon monoxide by calorimetry and compare it with the value calculated from spectroscopic data. The difference is the residual entropy of Section 5.2.
Pauling (1935) counts the arrangements of the hydrogen atoms in ice, the model of Example 2 of Section 5.2.
Haber (1920) gives his account of the synthesis of ammonia from nitrogen and hydrogen (Section 5.3).
McQuarrie, D. A., & Simon, J. D. (1997). Physical Chemistry: A Molecular Approach. University Science Books.
Dill, K. A., & Bromberg, S. (2011). Molecular Driving Forces: Statistical Thermodynamics in Biology, Chemistry, Physics, and Nanoscience (2nd ed.). Garland Science.
Hill, T. L. (1986). An Introduction to Statistical Thermodynamics. Dover Publications.
Clayton, J. O., & Giauque, W. F. (1932). The Heat Capacity and Entropy of Carbon Monoxide. Heat of Vaporization. Vapor Pressures of Solid and Liquid. Free Energy to 5000 K from Spectroscopic Data. Journal of the American Chemical Society, 54(7), 2610–2626. 10.1021/ja01346a004
Pauling, L. (1935). The Structure and Entropy of Ice and of Other Crystals with Some Randomness of Atomic Arrangement. Journal of the American Chemical Society, 57(12), 2680–2684. 10.1021/ja01315a102