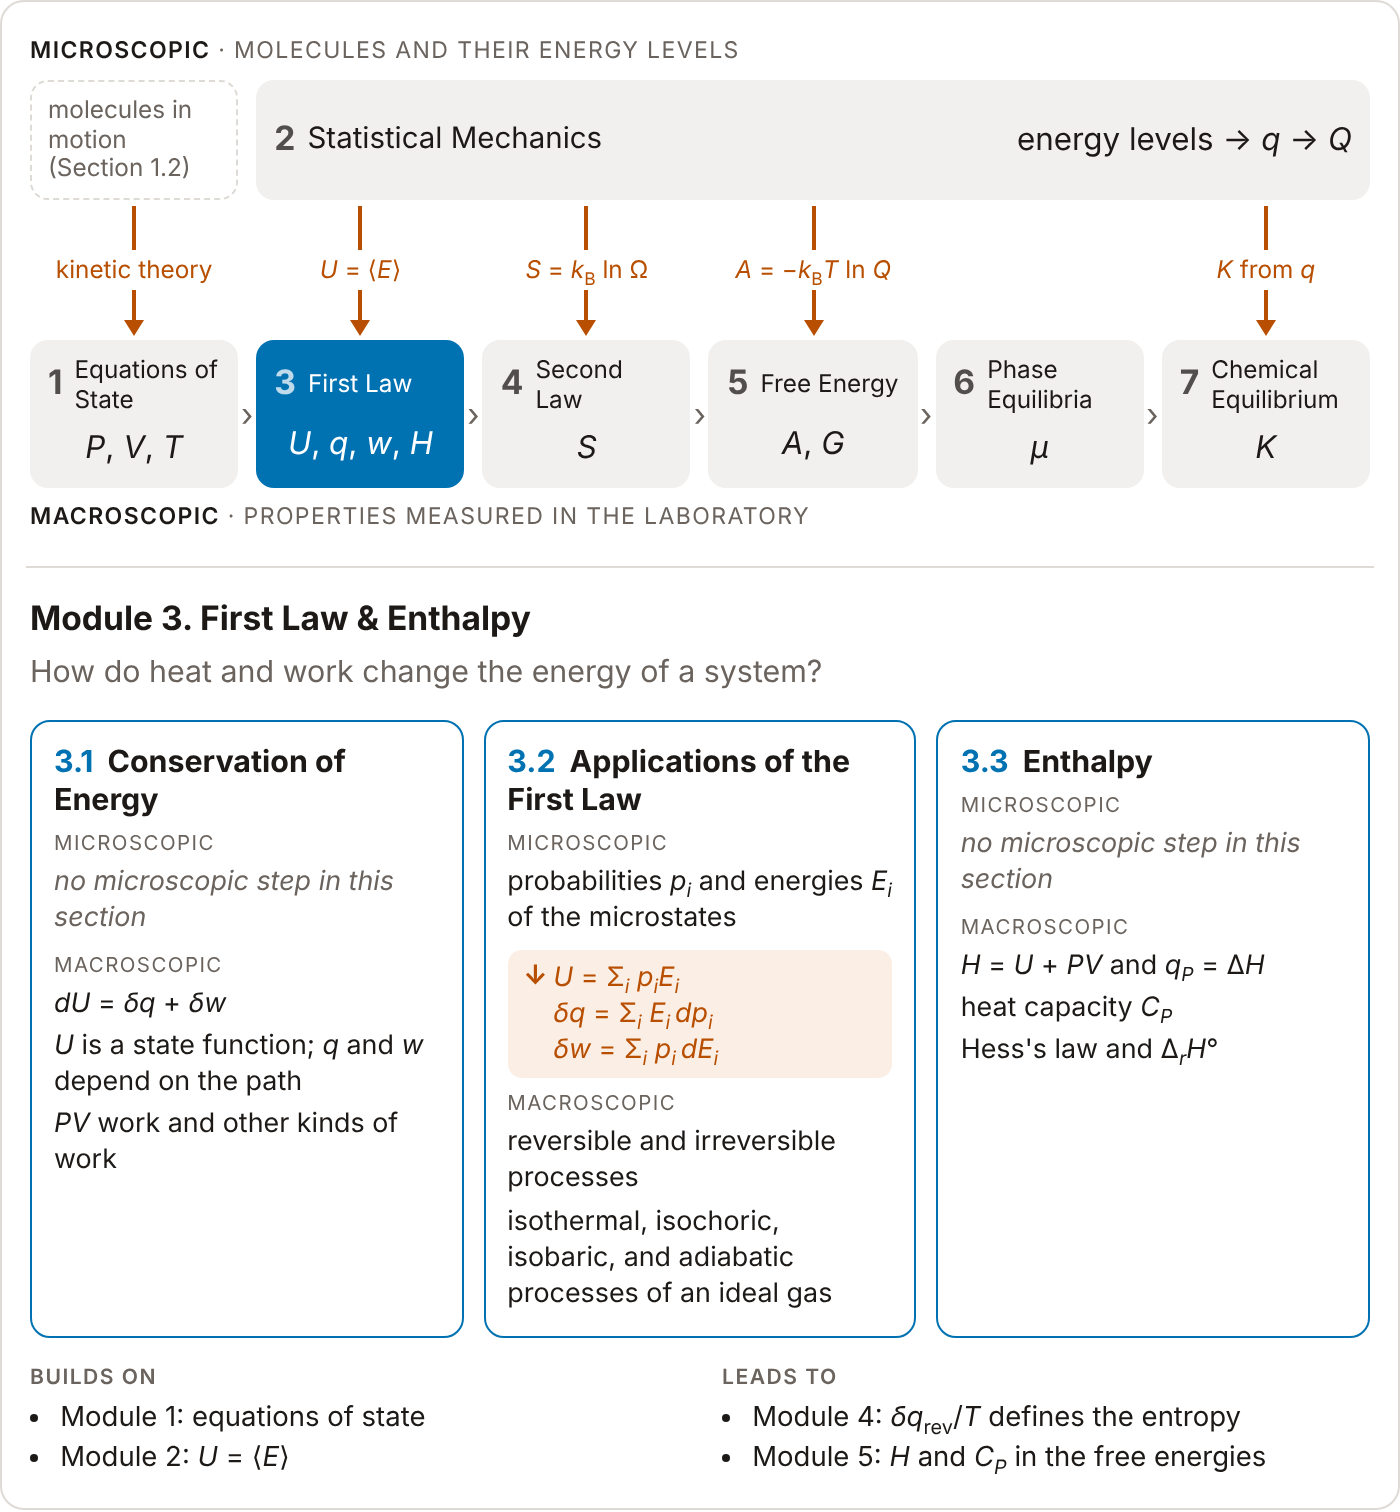
The course map shows where this module sits in the course and which relations connect the microscopic and macroscopic descriptions. Select a module to unfold its sections.
Joule found that a definite quantity of mechanical work always produces the same quantity of heat: about 4.19J of work raises the temperature of 1g of water by 1°C.
First Law of Thermodynamics
The internal energy change of a closed system equals the heat absorbed plus the work done on it:
In differential form: dU=δq+δw, or equivalently δq=dU−δw.
State Functions vs. Path Functions
U is a state function: ΔU depends only on the initial and final states, and dU is an exact differential (∮dU=0).
q and w are path functions: their values depend on the process path, and δq, δw are inexact differentials.
Sign Convention (Chemistry Convention)
q>0: heat absorbed by the system.
w>0: work done on the system.
Some textbooks define w as the work done by the system, so the convention of a source should be checked before one of its formulas is used.
For PV work: δw=−PextdV, so compression (dV<0) gives w>0. For a reversible process, Pext=P.
Generalized Work
Many work modes share a generalized force × generalized displacement structure: PV work (−PextdV), surface work (γdA), elastic work (fdl), electrical work (Edqel), and chemical work (μdN).
Free Expansion
Expansion into vacuum: Pext=0, so w=0. If the container is also insulated, q=0 and ΔU=0. For an ideal gas, ΔT=0 as well.
Free expansion is the extreme case of an irreversible process: the gas changes state while doing no work and exchanging no heat.
Quasi-static: carried out infinitesimally slowly; the system remains near equilibrium at all times.
Reversible: quasi-static and free of dissipative effects (friction, turbulence, viscous drag, etc.); can be reversed with no net change to system or surroundings.
Irreversible: any real process that violates one or more conditions for reversibility.
Reversible Work Bounds
For a given expansion (ΔV>0), a reversible process does the maximum work on the surroundings (∣wrev∣≥∣wirrev∣).
For a given compression, a reversible process requires the minimum work input.
Five-Step First Law Workflow
Choose two independent variables (e.g., V,T or P,T).
Rewrite the First Law in terms of those variables.
Apply process constraints (isothermal, isochoric, isobaric, adiabatic).
Specify the equation of state and an expression for U (e.g., ideal gas).
Compare with the heat capacity at constant volume, CV=(∂T∂U)V (Section 3.2).
Condition for qP=ΔH
The relation holds only when PV work is the sole form of work. The table of work terms in Section 3.1 lists other kinds of work.
Measuring Enthalpy Changes
For a change of temperature at constant pressure with no reaction or phase change, ΔH=∫TiTfCPdT≈CPΔT.
Calorimetry: a process exchanges heat with a known mass m of water (heat capacity CP per gram), whose temperature changes by ΔT. The heat absorbed by the process at constant pressure is qP=−mCPΔT=ΔH.
Standard States and Formation Enthalpies
Standard pressure: P∘=1bar; reference temperature typically 298.15 K.
The standard state of an element is its most stable form at P∘. Phosphorus is the usual exception: its reference state is white phosphorus.
Standard enthalpy of reaction: ΔrH∘=∑pνpHp∘−∑rνrHr∘, where Hp∘ and Hr∘ are standard molar enthalpies. Only differences of enthalpy can be measured.
Standard enthalpy of formation, ΔfH∘: enthalpy change when 1 mol of a compound in its standard state is formed from its elements in their standard states.
By convention, ΔfH∘=0 for elements in their standard states.
Hess’s Law
Because H is a state function, enthalpy changes are path-independent and additive:
Applicability: any expansion or compression. For reversible processes, Pext=P (system pressure). For irreversible processes, Pext is determined by the surroundings.
C. First Law in (V,T) Variables (for a reversible process with PV-only work)
One mole of nitrogen, treated as an ideal gas with CV,m=25R, starts in state A at TA=300K and PA=1.00bar. It ends in state B at PB=3.00bar with half the initial volume, VB=VA/2. Two reversible paths connect the states:
Path 1. The gas is compressed at constant pressure PA from VA to VB and then heated at constant volume VB until the pressure is PB.
Path 2. The gas is heated at constant volume VA until the pressure is PB and then compressed at constant pressure PB from VA to VB.
Use R=8.3145Jmol−1K−1.
(a) Find the temperature of state B and of the intermediate state of each path.
(b) Find w, ΔU, and q for each step of Path 1.
(c) Find w, ΔU, and q for each step of Path 2.
(d) Add the steps to obtain w, q, and ΔU for each path, in joules. Then calculate ΔU from the temperatures of A and B alone.
Solution
Every answer is written first as a multiple of nRTA=(1.00mol)(8.3145Jmol−1K−1)(300K)=2494J.
Temperatures
For a fixed amount of an ideal gas, T is proportional to PV. State B has three times the pressure and half the volume of state A, so TB=23TA=450K. The intermediate state of Path 1 has PA and VA/2, so T1=21TA=150K. The intermediate state of Path 2 has 3PA and VA, so T2=3TA=900K.
Relations for the two kinds of step
The work done on the gas in a reversible step is w=−∫PdV, and for an ideal gas ΔU=CVΔT=25nRΔT for any step. The heat then follows from the First Law, q=ΔU−w.
At constant volume, w=0 and q=ΔU.
At constant pressure, w=−PΔV=−nRΔT.
Path 1, compression at constant pressure (300K→150K)
the value obtained by adding q and w along either path.
Result.TB=450K, with intermediate temperatures of 150K (Path 1) and 900K (Path 2). Path 1 has w=+1247J and q=+1871J; Path 2 has w=+3742J and q=−624J. Both give ΔU=+3118J.
Problem 2. Adiabatic compression in a diesel cylinder
In the compression stroke of a diesel engine, the air in the cylinder is compressed to 1/18 of its initial volume before the fuel is injected. Treat the air as a diatomic ideal gas with CV,m=25R, and treat the compression as reversible and adiabatic. The air starts at T1=300K and P1=1.00bar. Use R=8.3145Jmol−1K−1.
(a) Follow the workflow of Section 3.2 to show that TV2/5 is constant along the path.
(b) Find the final temperature T2 and the final pressure P2.
(c) Find w, ΔU, and ΔH per mole of air.
(d) Diesel fuel ignites in hot air without a spark at about 500K. Does the compression alone bring the air to this temperature?
Solution
First Law for an adiabatic step
For an ideal gas in a reversible process with only PV work, δq=CVdT+PdV. Setting δq=0 and using P=nRT/V and CV=25nR,
The air reaches 953K, well above 500K, so fuel injected at the end of the stroke ignites without a spark.
Result.T2=953K and P2=57.2bar. Per mole of air, w=ΔU=+13.58kJ and ΔH=+19.01kJ. The compression heats the air past the ignition temperature of the fuel.
Problem 3. A reaction enthalpy at the process temperature
The table lists data from the NIST-JANAF tables. Treat each heat capacity as constant between 298.15K and 700K at its value at 500K.
Species
ΔfH∘(298.15K) (kJmol−1)
CP,m(500K) (Jmol−1K−1)
ΔfH∘(700K) (kJmol−1)
SO2(g)
−296.842
46.576
−306.291
O2(g)
0
31.091
0
SO3(g)
−395.765
63.100
−405.014
(a) Find ΔrH∘ at 298.15K.
(b) Find ΔrH∘ at 700K from a cycle of three steps at constant pressure: cool the reactants from 700K to 298.15K, carry out the reaction at 298.15K, and heat the product to 700K.
(c) Compare your result with the value that follows from the formation enthalpies at 700K.
(d) How much heat is released when 1.00kg of SO3 (M=80.063gmol−1) forms at 700K and constant pressure?
The reactants are one mole of SO2 and half a mole of O2, with a combined heat capacity of 46.576+21(31.091)=62.122JK−1. With ΔT=298.15−700=−401.85K,
The formation enthalpies at 700K give ΔrH∘(700K)=(−405.014)−(−306.291)=−98.723kJmol−1. The estimate with constant heat capacities differs from it by 0.19kJmol−1, or 0.2%.
Heat released per kilogram
One kilogram of SO3 is 1000/80.063=12.49mol. At constant pressure the heat absorbed is qP=ΔH, so
The reaction releases 1.23MJ of heat per kilogram of SO3.
Result.ΔrH∘=−98.92kJmol−1 at 298.15K and −98.53kJmol−1 at 700K from the cycle, against the tabulated −98.72kJmol−1. Forming 1.00kg of SO3 at 700K releases 1.23MJ.
A bomb calorimeter holds its contents at constant volume. It is calibrated by burning 0.9000g of benzoic acid, whose energy of combustion at constant volume is −26.434kJg−1; the temperature of the calorimeter rises by 2.379K. Then 1.500g of urea (M=60.056gmol−1) is burned in the same calorimeter near 298K, and the temperature rises by 1.582K. The combustion reaction is
From H=U+PV, ΔH=ΔU+Δ(PV). With ideal gases and negligible volumes of the solid and the liquid, Δ(PV)=ΔngasRT at constant temperature. The reaction consumes 23 mol of gas and forms 2 mol, so Δngas=+21 and
McQuarrie & Simon (1997), Chapter 19 (the first law of thermodynamics).
Dill & Bromberg (2011), Chapter 3 (heat, work, and energy) and Chapter 7 (the logic of thermodynamics).
Original papers
Joule (1850) reports the paddle-wheel experiments of Section 3.1 and the mechanical equivalent of heat obtained from them.
Hess (1840) states that the heat of a chemical change is the same whether the change occurs in one step or in several, which is Hess’s law of Section 3.3.
McQuarrie, D. A., & Simon, J. D. (1997). Physical Chemistry: A Molecular Approach. University Science Books.
Dill, K. A., & Bromberg, S. (2011). Molecular Driving Forces: Statistical Thermodynamics in Biology, Chemistry, Physics, and Nanoscience (2nd ed.). Garland Science.
Joule, J. P. (1850). On the Mechanical Equivalent of Heat. Philosophical Transactions of the Royal Society of London, 140, 61–82. 10.1098/rstl.1850.0004
Hess, H. (1840). Thermochemische Untersuchungen. Annalen Der Physik, 126(6), 385–404. 10.1002/andp.18401260620