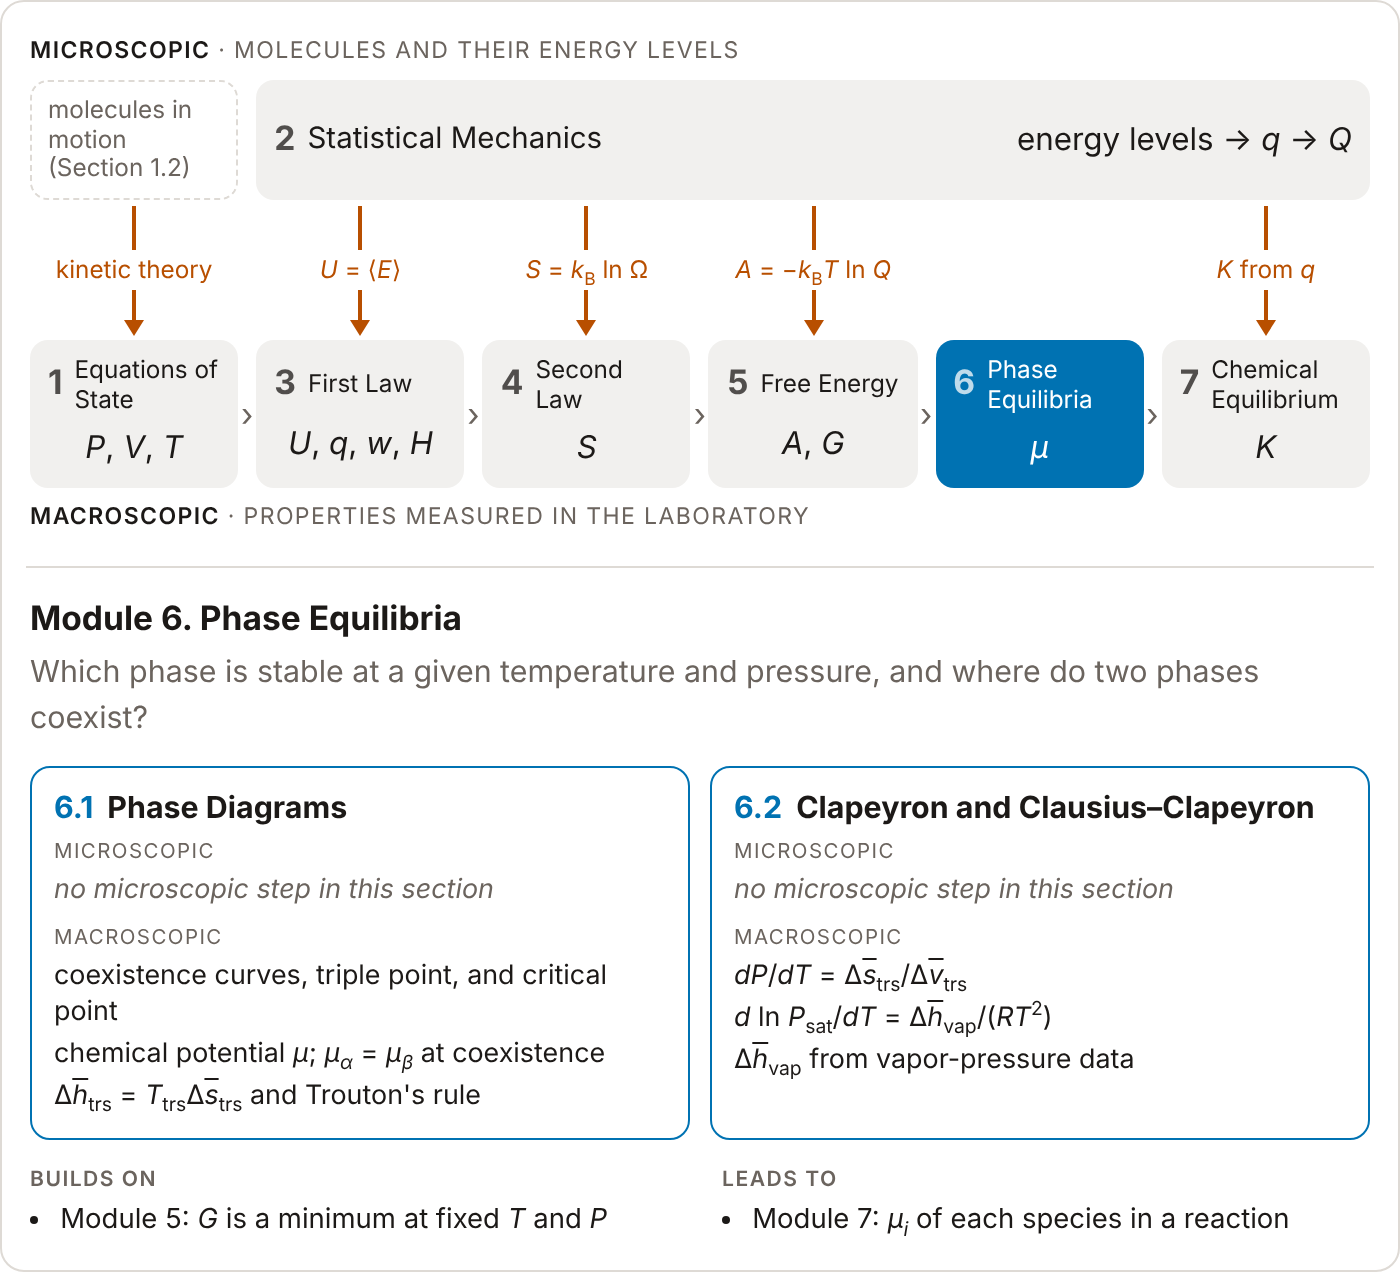
The course map shows where this module sits in the course and which relations connect the microscopic and macroscopic descriptions. Select a module to unfold its sections.
Single-phase regions (solid, liquid, gas) are separated by coexistence curves on which two phases are simultaneously in equilibrium.
Triple point: the unique (Ttp,Ptp) at which solid, liquid, and gas coexist.
Critical point: the point at which the liquid–vapor coexistence curve terminates; beyond it, liquid and gas become indistinguishable (a supercritical fluid).
The solid–liquid line does not terminate in a critical point because solid and liquid differ qualitatively (long-range translational order), while liquid and gas differ only quantitatively.
Open Systems and the Chemical Potential
A closed system cannot describe two coexisting phases by itself; each phase must be treated as an open system whose amount of matter can change.
This is the Euler relation of Section 5.1, G=μN, written per mole.
Phase-Equilibrium Condition
Letting the amount of phase α change by dnα at fixed T and P, with dnβ=−dnα, gives dG=(μα−μβ)dnα. Requiring dG=0 for an arbitrary small dnα yields
Physically: matter flows from higher μ to lower μ and stops when they are equal. Because this is a single scalar constraint on two variables, it picks out a curve in the (T,P) plane — the coexistence curve.
Graphical View: gˉ vs. T at Fixed P
From dG=−SdT+VdP: (∂gˉ/∂T)P=−sˉ.
Every phase has negative slope; the larger sˉ, the steeper (more negative) the slope.
Since sˉg>sˉl>sˉs, the gas line is steepest, then liquid, then solid. Each intersection marks a first-order transition at which the stable (lowest-gˉ) phase changes.
Graphical View: gˉ vs. P at Fixed T
From dG=−SdT+VdP: (∂gˉ/∂P)T=vˉ.
Every phase has positive slope; the larger vˉ, the steeper the slope.
Since vˉg≫vˉl≳vˉs, compressing the system favors condensed phases over gas.
Water anomaly:vˉs>vˉl (ice is less dense than liquid water). Concept Check 5 of Section 6.1 asks what follows for the two curves.
Latent Heat and Entropy Jump at a First-Order Transition
A first-order transition is one at which the first derivatives of G (S and V) are discontinuous across the coexistence curve.
At coexistence, μα=μβ gives Δgˉtrs=0; combined with Δgˉ=Δhˉ−TΔsˉ:
Approximate constancy reflects that the entropy gained on vaporization is dominated by the phase change itself (volume increase, loss of short-range order) rather than the liquid’s chemical identity.
Hydrogen-bonded liquids (water, the lower alcohols, ammonia) lie well above the Trouton value. For water, Δsˉvap≈109Jmol−1K−1, roughly 25% above Trouton. Liquids that boil at very low temperatures (neon, hydrogen, helium) lie well below it.
Exact for any first-order transition between pure phases.
Sign and Magnitude of dP/dT
The sign is set by Δvˉ, since Δhˉ and T are positive at any “normal” transition. Most substances have vˉl>vˉs (positive melting slope) and vˉg≫vˉl (positive boiling slope).
The magnitude is set by Δvˉ: solid–liquid transitions have very small Δvˉ and therefore nearly vertical coexistence curves; liquid–vapor transitions have very large Δvˉ and shallow coexistence curves.
This is why melting points are nearly pressure-independent at ordinary pressures while boiling points shift strongly.
Water: Negative-Slope Ice–Water Line
Because vˉs>vˉl for water, Δvˉfus<0 and therefore dP/dT<0 along the ice–water line. Numerically, dP/dT≈−135barK−1, equivalently dT/dP≈−7.4mKbar−1.
The effect is far too small to explain ice skating by “pressure melting”; surface friction and a pre-existing liquid layer dominate.
Liquid–Vapor Approximation
Two approximations collapse the Clapeyron equation into a far more useful form for the liquid–vapor line:
vˉg≫vˉl, so Δvˉvap≈vˉg (for water at 1 atm the ratio exceeds 103).
The vapor is treated as an ideal gas, vˉg=RT/Psat.
Two-point form: two (T,P) points give Δhˉvap directly.
Linearized form: a plot of lnPsat vs. 1/T should be a straight line with slope −Δhˉvap/R, if the constant-Δhˉvap approximation holds.
Diagnosing Breakdown and the Local Slope
A lnP-vs-1/T plot can look straight even when Δhˉvap is varying. A residual plot is the reliable diagnostic: systematic curvature (not random scatter) indicates breakdown of the constant-Δhˉvap assumption.
The local slope from finite differences obeys the exact relation
Where the vapor is nearly ideal (ΔZ≈1), the local slope estimates Δhˉvap(T); it exceeds Δhˉvap as the vapor becomes denser (by 8 % for water at 180 °C).
Δhˉvap(T) decreases with rising T. At the critical point, where the liquid and vapor phases merge, Δhˉvap and Δvˉvap vanish together.
Molecular Reading of Δhˉvap
For water, Δhˉvap≈40kJmol−1 is consistent with breaking roughly two hydrogen bonds per molecule (each worth ∼20kJmol−1) on going from liquid to vapor — the macroscopic shadow of the intermolecular forces of Section 1.4.
Applicability: two phases of a pure substance in equilibrium at fixed T and P. A single scalar constraint on two variables defines the coexistence curve in the (T,P) plane.
Applicability: pure substance, single phase. Used graphically to identify the stable phase (lowest gˉ) and to read off entropy and molar volume from slopes.
F. Latent Heat / Entropy Jump at a First-Order Transition
Applicability: empirical benchmark at the normal boiling point for ordinary (non-hydrogen-bonded, non-associated) liquids. Hydrogen-bonded liquids lie well above it, and liquids that boil at very low temperatures lie well below it.
Applicability: exact for any first-order transition between pure phases. The sign of dP/dT is set by Δvˉ; the magnitude is dominated by ∣Δvˉ∣ (small for solid–liquid, large for liquid–vapor).
I. Clausius–Clapeyron Equation (Differential Form)
Applicability: liquid–vapor coexistence with Δhˉvap approximately constant over the interval [T1,T2]. A plot of lnPsat vs. 1/T is a straight line with slope −Δhˉvap/R.
Applicability: exact along the liquid–vapor coexistence curve. Where the vapor is nearly ideal (ΔZ≈1), the local slope from finite differences of vapor-pressure data estimates the temperature-dependent Δhˉvap(T). As T→Tc, Δhˉvap→0 and Δvˉvap→0 together.
Toluene boils at Tb=383.75K under 1atm, where Δhˉvap=33.23kJmol−1. At that temperature the molar heat capacities are cˉP=184.6Jmol−1K−1 for the liquid and 137.8Jmol−1K−1 for the vapor (NIST Chemistry WebBook). The pressure is 1atm throughout, and Δgˉvap=gˉg−gˉl=μg−μl.
(a) Find Δsˉvap at Tb, and compare it with the value of Trouton’s rule.
(b) Treat Δhˉvap and Δsˉvap as independent of temperature. Find Δgˉvap at 363.75K and at 403.75K, and name the stable phase at each temperature.
(c) Treat the two heat capacities as constant instead. Find Δhˉvap, Δsˉvap, and Δgˉvap at the same two temperatures. By how much does each Δgˉvap differ from the result of part (b)?
Solution
Entropy of vaporization
At the boiling point the two phases coexist, so Δgˉvap=Δhˉvap−TbΔsˉvap=0 and
At 363.75K, μg>μl: vapor at 1atm would condense spontaneously, and the liquid is the stable phase. At 403.75K, μg<μl, and the vapor is the stable phase.
Constant heat capacities
Take the liquid from T to Tb, vaporize it there, and bring the vapor back to T. With ΔcˉP=cˉP,g−cˉP,l=137.8−184.6=−46.8Jmol−1K−1, the heating integrals of Section 6.1 give
Each Δgˉvap moves by about +25Jmol−1, which is 1.4% of its size. The enthalpy term and the entropy term each change by about 0.9kJmol−1, and the two changes nearly cancel in Δgˉvap. The signs, and with them the stable phases, are the same as in part (b).
Result.Δsˉvap=86.59Jmol−1K−1 at the boiling point. Δgˉvap is +1.73kJmol−1 at 363.75K (liquid stable) and −1.73kJmol−1 at 403.75K (vapor stable). The heat capacities change these values to +1.76kJmol−1 and −1.71kJmol−1.
Problem 2. Latent heats of ammonia from its phase diagram
The table lists two points on the liquid–vapor coexistence curve of ammonia, with the tabulated enthalpy of vaporization at each (NIST Chemistry WebBook).
Point
T (K)
Psat (kPa)
Δhˉvap (kJmol−1)
Triple point
195.50
6.056
25.36
Normal boiling point
239.83
101.325
23.33
Just below the triple point, the enthalpy of sublimation is Δhˉsub=31.2kJmol−1. Use R=8.3145Jmol−1K−1.
(a) Estimate Δhˉvap from the two pressures with the integrated Clausius–Clapeyron equation, and compare the estimate with the tabulated values.
(b) Find Δhˉfus at the triple point from Δhˉsub and the tabulated Δhˉvap.
(c) Find the slopes dP/dT of the sublimation curve and of the vaporization curve at the triple point. Treat the vapor as an ideal gas and neglect the molar volumes of the solid and the liquid.
(d) Near the triple point, the melting curve of ammonia follows dT/dP=aT with a=4×10⁻⁵atm−1Haar & Gallagher, 1977. Find dP/dT of the melting curve at the triple point in PaK−1, and compare it with the slopes of part (c). Then find Δvˉfus from the Clapeyron equation and the result of part (b).
Solution
Enthalpy of vaporization from two points
Let point 1 be the triple point and point 2 the normal boiling point. The integrated Clausius–Clapeyron equation, solved for the enthalpy, is
The estimate lies between the tabulated values at the two ends of the interval, 25.36 and 23.33kJmol−1.
Enthalpy of fusion
At the triple point the solid, the liquid, and the vapor coexist at one temperature and one pressure. Enthalpy is a state function, so subliming the solid changes the enthalpy by the same amount as melting it and then vaporizing the liquid:
With Δvˉ≈vˉg=RT/P, the Clapeyron equation becomes dP/dT=PΔhˉ/(RT2) for either curve. At the triple point, RT2=(8.3145)(195.50)2=3.1778×10⁵JKmol−1 and P=6056Pa, so
Result.Δhˉvap≈24.78kJmol−1 from the two pressures, and Δhˉfus=5.8kJmol−1. At the triple point the slopes are 595PaK−1 (sublimation), 483PaK−1 (vaporization), and 1.3×10⁷PaK−1 (melting), with Δvˉfus=2.3cm3mol−1.
Problem 3. A sublimation curve and the triple-point pressure
Sulfur hexafluoride, the insulating gas of high-voltage switchgear, sublimes at atmospheric pressure: the solid and the vapor coexist at 209.3K and 1.01325bar. Its enthalpy of sublimation is Δhˉsub=23.3kJmol−1, and its triple point lies at Ttp=223.56K (NIST Chemistry WebBook). Treat the vapor as an ideal gas, neglect the molar volume of the solid, and take Δhˉsub as constant. Use R=8.3145Jmol−1K−1.
(a) Start from the Clapeyron equation and show that the sublimation curve obeys the Clausius–Clapeyron equation with Δhˉsub in place of Δhˉvap. Find the slope dP/dT of the sublimation curve at 1.01325bar.
(b) Estimate the triple-point pressure Ptp, and compare it with the measured 2.314bar.
(c) Can liquid sulfur hexafluoride be a stable phase at 1.01325bar? Give the reason.
Solution
Clausius–Clapeyron equation for sublimation
The Clapeyron equation holds for any first-order transition, so for solid to vapor
No. On the phase diagram the liquid region begins at the triple point, so the liquid is a stable phase only at pressures above Ptp=2.314bar. Atmospheric pressure lies below Ptp. A path of rising temperature at 1.01325bar crosses only the sublimation curve, at 209.3K, and the solid passes directly into the vapor.
Result.dP/dT=6.48kPaK−1 at 1.01325bar, and Ptp≈2.38bar against the measured 2.314bar. Liquid sulfur hexafluoride is not stable at atmospheric pressure.
Problem 4. Vapor pressure of propane from an equation
with A=3.98292, B=819.296K, and C=−24.417K. Use R=8.3145Jmol−1K−1=0.083145Lbarmol−1K−1.
(a) Find the normal boiling point, where Psat=1.01325bar, from the equation. Compare it with the measured 231.04K.
(b) Differentiate the equation to find dlnPsat/dT. Use the Clausius–Clapeyron equation to estimate Δhˉvap at 231.04K, and compare the estimate with the calorimetric 18.77kJmol−1.
(c) At the normal boiling point, vˉg=18.25Lmol−1 and vˉl=0.076Lmol−1. Find ΔZ, and repeat the estimate of part (b) with the exact relation between the slope and Δhˉvap.
(d) A cylinder that holds liquid propane and its vapor stands in the sun at 318.15K (45∘C). Find the pressure in the cylinder, and compare it with 15.34bar from the NIST Chemistry WebBook.
Solution
Normal boiling point
Solve the equation for the temperature. With log10(1.01325)=0.00572,
Result. The equation gives a normal boiling point of 230.41K and Psat=15.62bar at 318.15K. At the boiling point, Δhˉvap=19.61kJmol−1 from the Clausius–Clapeyron equation and 18.80kJmol−1 with ΔZ=0.9586, against the calorimetric 18.77kJmol−1.
McQuarrie & Simon (1997), Chapter 23, Sections 23.1 to 23.5 (phase equilibria).
Dill & Bromberg (2011), Chapter 14 (equilibria between liquids, solids, and gases) and Chapter 25 (phase transitions).
Original papers
Clapeyron (1834) relates the slope of the vapor-pressure curve to the heat of vaporization, the first form of the Clapeyron equation of Section 6.2. The English translation by R. Taylor is in Scientific Memoirs1, 347–376 (1837).
Clausius (1850) rederives Clapeyron’s relation from the equivalence of heat and work (Section 6.2). An English translation appeared in Philosophical Magazine2, 1–21 and 102–119 (1851).
Trouton (1884) observes that the molar heat of vaporization divided by the boiling temperature is nearly the same for many liquids, which is Trouton’s rule of Section 6.1.
Haar, L., & Gallagher, J. S. (1977). The Equation of State for Ammonia (Techreport NBSIR 77-1409). National Bureau of Standards. 10.6028/NBS.IR.77-1409
McQuarrie, D. A., & Simon, J. D. (1997). Physical Chemistry: A Molecular Approach. University Science Books.
Dill, K. A., & Bromberg, S. (2011). Molecular Driving Forces: Statistical Thermodynamics in Biology, Chemistry, Physics, and Nanoscience (2nd ed.). Garland Science.
Clapeyron, É. (1834). Mémoire sur la puissance motrice de la chaleur. Journal de l’École Polytechnique, 14, 153–190.
Clausius, R. (1850). Ueber die bewegende Kraft der Wärme und die Gesetze, welche sich daraus für die Wärmelehre selbst ableiten lassen. Annalen Der Physik, 155(3), 368–397. 10.1002/andp.18501550306
Trouton, F. (1884). On Molecular Latent Heat. The London, Edinburgh, and Dublin Philosophical Magazine and Journal of Science, 18(110), 54–57. 10.1080/14786448408627563